
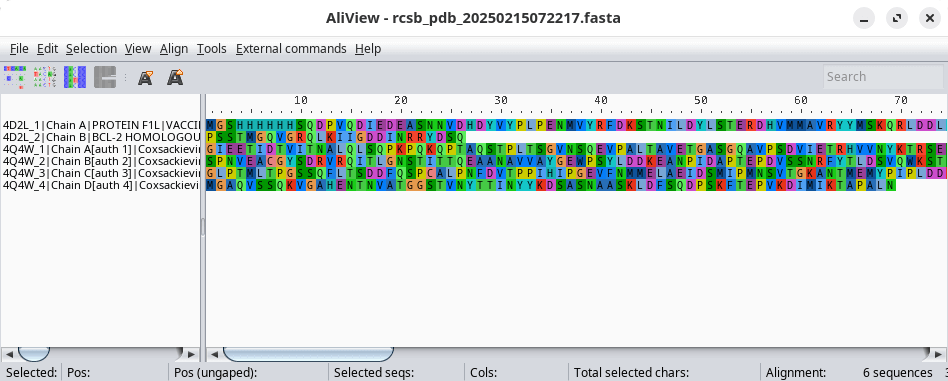
AliView is an alignment viewer and editor of DNA / aminoacid sequences. It aims to be one of the fastest and most intuitive to use.
The general idea when designing this program has always been usability and speed, all new functions are optimized so they do not affect the general performance and capability to work swiftly with large alignments.
A need to easily sort, view, remove, edit and merge sequences from large transcriptome datasets initiated the work with the program.
Key Features
- Supports Fasta, Nexus, Phylip, Clustal or MSF-format (unlimited file sizes).
edit (manually). - Align, add and align automatically (with MUSCLE or MAFFT or any other aligner of your choice).
- Find degenerate primers in conserved regions in an alignment of mixed species.
- On the fly translation of nucleotides to amino acids.
- Various visual cues to highlight consensus characters or characters deviating from the consensus.
- Simple copy/paste/drop/remove of sequences/files.
- Very simple to use “external interface” that lets you invoke your other favorite programs (you could for example automatically have the alignment sent to FastTree and then automatically opened in FigTree).
- Cross-platform support – runs under Linux, macOS, and Windows.
Website: www.ormbunkar.se/aliview
Support: GitHub Code Repository
Developer: Department of Systematic Biology (Uppsala University)
License: GNU General Public License v3.0
AliView is written in Java. Learn Java with our recommended free books and free tutorials.
Related Software
| Bioinformatics Tools | |
|---|---|
| Bioconductor | Analysis and comprehension of high-throughput genomic data |
| Biopython | Tools for biological computation written in Python |
| UGENE | Set of integrated bioinformatics software |
| BioPerl | Perl tools for computational molecular biology |
| GROMACS | Versatile package to perform molecular dynamics |
| IGV | High-performance visualization genome browser tool |
| GATK | Genomic analysis toolkit focused on variant discovery |
| BioJava | Provides Java tools for processing biological data |
| InterMine | Integrate biological data sources |
| bedtools | Powerful toolset for genome arithmetic |
| EMBOSS | The European Molecular Biology Open Software Suite |
| BLAST | Algorithm for comparing primary biological sequence information |
| Galaxy | Web-based platform for data-intensive computational research |
| minimap2 | Versatile sequence alignment program |
| Scanpy | Scalable analysis of single-cell gene expression data |
| Jalview | Multiple sequence alignment editing, visualisation and analysis |
| SeqKit | Fast toolkit for manipulating FASTA and FASTQ sequence files |
| samtools | Manipulate next-generation sequencing data |
| MultiQC | Aggregates results from multiple bioinformatics analyses |
| BCFtools | Variant calling and manipulating files in the Variant Call Format |
| Foldseek | Fast searching and comparison of large protein structure datasets |
| FastQC | Quality control tool for high throughput sequence data |
| SPAdes | Versatile toolkit for assembling and analysing sequencing data |
| GenomeTools | Collection of bioinformatics tools |
| AliView | Alignment viewer and editor |
| mothur | Analyze microbial communities |
| Bandage | Visualising de novo assembly graphs |
| cramino | BAM/CRAM quality evaluation |
| abPOA | Adaptive banded Partial Order Alignment |
Read our verdict in the software roundup.
 Explore our carefully curated directory of recommended free and open source software, covering every major software category. The directory forms part of our extensive collection of articles for Linux enthusiasts. It includes hundreds of detailed reviews, together with free and open source alternatives to proprietary software from companies such as Google, Microsoft, Apple, Adobe, IBM, Cisco, Oracle, and Autodesk. LinuxLinks also covers interesting projects worth exploring, Linux-compatible hardware, free programming books and tutorials, and much more. Know a useful free and open source Linux application that we haven’t covered? Tell us about it using our submission form. |
Please read our Comment Policy before commenting.